Atomaktin Capsule
PDF Leaflet Revision Date: 21 August 2025
Clinical Summary
Quick overview from the medicine insert
Indication
Treatment of ADHD in children, adolescents, and adults.
Dosage (summary)
Children <70 kg: 0.5 mg/kg/day; Adults: 40 mg initial, 80 mg maintenance.
Special Populations
- Hepatic impairment
- End-stage renal disease
Pregnancy & Breastfeeding
Not recommended during pregnancy or breastfeeding.
Key Drug Interactions
- MAOIs
- CYP2D6 inhibitors
- Salbutamol
Contraindications
- Hypersensitivity
- Uncontrolled hypertension
- Narrow angle glaucoma
- Severe cardiovascular disorders
Common side effects
- Headache
- Abdominal pain
- Decreased appetite
- Nausea
- Fatigue
Counselling Points
- Monitor for suicidal thoughts
- Avoid in severe cardiac conditions
- Use caution when driving or operating machinery
Serious warnings
- Increased risk of suicidal ideation in children and adolescents
- Cardiovascular effects
Important Disclaimer
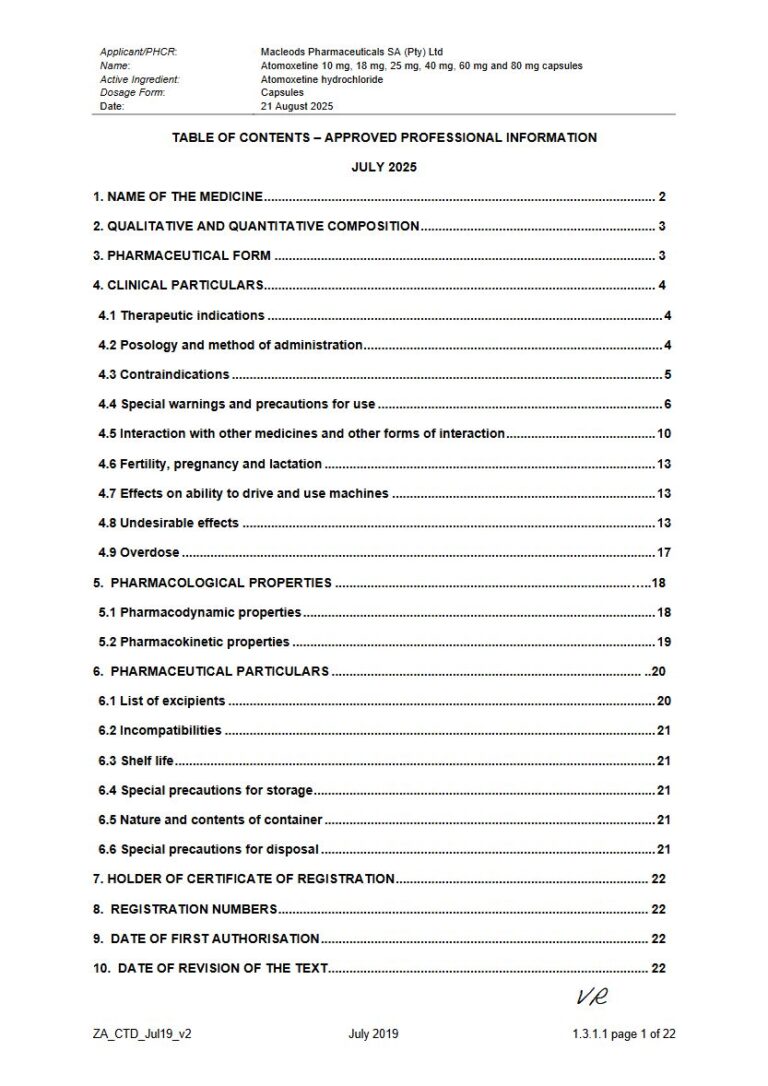
The Atomaktin Capsule professional information leaflet below is the property of Macleods Pharmaceuticals Sa and is provided on Medinsert exactly as issued, with no.. alterations or editorial changes. We make every effort to keep content current by updating documents as soon as new versions become available. Medinsert serves as a trusted access point for healthcare professionals, but does not replace official sources or clinical judgement. For more details, please read our full disclaimer. read more>>
 Healthcare Professionals Only
This content is for registered healthcare professionals
Sign in or create a free account to read the full package insert.
Free for HPCSA-registered professionals. Powered by Medinsert.
Clinical Particulars
Section 4 of the official insert — extracted exactly as issued, no alterations
4.1 Therapeutic indications
ATOMAKTIN is indicated for the treatment of Attention - Deficit/Hyperactivity Disorder (ADHD) in children 6 years of age or older, adolescents and adults.
4.2 Posology and method of administration
Posology Treatment must be initiated by or under the supervision of a medical practitioner with appropriate knowledge and experience of childhood and/or adolescent behavioural disorders (for example, paediatrician or child/adolescent psychiatrist) (See section 4.4). The recommended initial dose and subsequent dosage escalations of ATOMAKTIN should not be exceeded because of potential side effects (see section 4.8). ATOMAKTIN capsules are not intended to be opened. ATOMAKTIN is an ocular irritant. In the event of capsule content coming into contact with the eye, the affected eye should be flushed immediately with water, and medical advice obtained. Hands and any potentially contaminated surfaces should be washed as soon as possible.
Dosing of children and adolescents up to 70 kg body weight: ATOMAKTIN should be initiated at a total daily dose of approximately 0,5 mg/kg. The initial dose should be maintained for a minimum of 7 days prior to upward dose titration according to clinical response and tolerability. The recommended maintenance dose is approximately 1,2 mg/kg/day (depending on the patientu2019s weight and available dosage strengths of ATOMAKTIN). No additional benefit has been demonstrated for doses higher than 1,2 mg/kg/day.
Dosing of children and adolescents over 70 kg body weight and adults: ATOMAKTIN should be initiated at a total daily dose of 40 mg. The initial dose should be maintained for a minimum of 7 days prior to upward dose titration according to clinical response and tolerability. The recommended maintenance dose is 80 mg. No additional benefit has been demonstrated for doses higher than 80 mg. The maximum recommended total daily dose for adults is 80 mg.
Special populations For those ADHD patients who have hepatic insufficiency or end-stage renal disease, cautious titration of ATOMAKTIN to the desired clinical response is recommended. ATOMAKTIN clearance may be reduced in patients with hepatic insufficiency. ATOMAKTIN may exacerbate hypertension in patients with end-stage renal disease. ATOMAKTIN may be discontinued without tapering the dose. Long term use: No fixed dose-response studies have been conducted in adults. The recommended daily dose of 80 mg reflects the optimal daily dose of 1,2 mg/kg/day demonstrated in children and adolescents. No controlled long-term studies have been conducted in adults. Open-label study data from 384 patients with up to 97 weeks of treatment with atomoxetine are consistent with maintenance of efficacy in long-term treatment.
Method of administration For oral use. ATOMAKTIN may be taken with or without food.
4.3 Contraindications
ATOMAKTIN is contraindicated:
- Hypersensitivity to the active substance or to any of the excipients listed in section 6.1.
- ATOMAKTIN should not be used in patients with uncontrolled hypertension or impairment of liver function.
- Atomoxetine should not be used in combination with monoamine oxidase inhibitors (MAOI) including linezolid. Atomoxetine should not be used within a minimum of 2 weeks after discontinuing therapy with MAOI. Treatment with MAOI should not be initiated within 2 weeks after discontinuing atomoxetine.
- Atomoxetine should not be used in patients with narrow angle glaucoma, as in clinical trials the use of atomoxetine was associated with an increased incidence of mydriasis.
- Atomoxetine should not be used in patients with severe cardiovascular or cerebrovascular disorders (see section 4.4). Severe cardiovascular disorders may include severe hypertension, heart failure, arterial occlusive disease, angina, haemodynamically significant congenital heart disease, cardiomyopathies, myocardial infarction, potentially life-threatening dysrhythmias and channelopathies (disorders caused by the dysfunction of ion channels). Severe cerebrovascular disorders may include cerebral aneurysm or stroke.
- Atomoxetine should not be used in patients with pheochromocytoma or a history of pheochromocytoma (see section 4.4).
4.4 Special warnings and precautions for use
Suicide - related behaviour Suicide related behaviour (suicide attempts and suicidal ideation) has been reported in patients treated with atomoxetine. In double blind clinical trials, suicide related behaviours were uncommon but more frequently observed among children and adolescents treated with atomoxetine compared to those treated with placebo, where there were no events. In adult double-blind clinical trials there was no difference in the frequency of suicide related behaviour between atomoxetine and placebo. Patients who are being treated for ADHD should be carefully monitored for the appearance or worsening of suicide related behaviour.
Sudden death and pre-existing cardiac abnormalities Sudden death has been reported in patients with structural cardiac abnormalities who were taking atomoxetine at usual doses. Although some serious structural cardiac abnormalities alone carry an increased risk of sudden death, atomoxetine should only be used with caution in patients with known serious structural cardiac abnormalities and in consultation with a cardiac specialist.
Cardiovascular effects Atomoxetine can affect heart rate and blood pressure. Most patients taking atomoxetine experience a modest increase in heart rate (mean <10 bpm) and/or increase in blood pressure (mean <5 mm Hg) (see section 4.8). However, combined data from controlled and uncontrolled ADHD clinical trials show that approximately 8-12% of children and adolescents, and 6-10% adults experience more pronounced changes in heart rate (20 beats per minute or greater) and blood pressure (15-20 mmHg or greater). Analysis of these clinical trial data showed that approximately 15-26% of children and adolescents, and 27-32% of adults experiencing such changes in blood pressure and heart rate during atomoxetine treatment had sustained or progressive increases. Long-term sustained changes in blood pressure may potentially contribute to clinical consequences such as myocardial hypertrophy. As a result of these findings, patients who are being considered for treatment with ATOMAKTIN should have a careful history and physical exam to assess for the presence of cardiac disease, and should receive further specialist cardiac evaluation if initial findings suggest such history or disease. It is recommended that heart rate and blood pressure be measured and recorded before treatment is started and, during treatment, after each adjustment of dose and then at least every 6 months to detect possible clinically important increases. For paediatric patients the use of a centile chart is recommended. For adults, current reference guidelines for hypertension should be followed. ATOMAKTIN should not be used in patients with severe cardiovascular or cerebrovascular disorders (see section 4.3). ATOMAKTIN should be used with caution in patients whose underlying medical conditions could be worsened by increases in blood pressure and heart rate, such as patients with hypertension, tachycardia, or cardiovascular or cerebrovascular disease.
Patients who develop symptoms such as palpitations, exertional chest pain, unexplained syncope, dyspnoea or other symptoms suggestive of cardiac disease during atomoxetine treatment should undergo a prompt specialist cardiac evaluation. In addition, ATOMAKTIN should be used with caution in patients with congenital or acquired long QT or a family history of QT prolongation (see sections 4.5 and 4.8). As orthostatic hypotension has also been reported, ATOMAKTIN should be used with caution in any condition that may predispose patients to hypotension or conditions associated with abrupt heart rate or blood pressure changes. ATOMAKTIN should not be used in patients with Raynaudu2019s phenomenon.
Cerebrovascular effects Patients with additional risk factors for cerebrovascular conditions (such as a history of cardiovascular disease, concomitant medications that elevate blood pressure) should be assessed at every visit for neurological signs and symptoms after initiating treatment with atomoxetine.
Hepatic effects Spontaneous reports of liver injury, manifested by elevated hepatic enzymes and bilirubin with jaundice, have been reported. Severe liver injury, including acute liver failure, have been reported. ATOMAKTIN should be discontinued in patients with jaundice or laboratory evidence of liver injury, and should not be restarted.
Psychotic or manic symptoms Treatment emergent psychotic or manic symptoms, e.g., hallucinations, delusional thinking, mania or agitation in patients without a prior history of psychotic illness or mania can be caused by atomoxetine at usual doses. If such symptoms occur, consideration should be given to a possible causal role of atomoxetine, and discontinuation of treatment should be considered. The possibility that ATOMAKTIN will cause the exacerbation of pre-existing psychotic or manic symptoms cannot be excluded.
Aggressive behaviour, hostility or emotional lability Hostility (predominantly aggression, oppositional behaviour and anger) was more frequently observed in clinical trials among children, adolescents and adults treated with atomoxetine as in ATOMAKTIN compared to those treated with placebo. Emotional lability was more frequently observed in clinical trials among children treated with atomoxetine compared to those treated with placebo. Patients should be closely monitored for the appearance or worsening of aggressive behaviour, hostility or emotional lability.
Possible allergic events Allergic reactions, including anaphylactic reactions, rash, angioneurotic oedema, and urticaria, have been reported in patients taking atomoxetine.
Seizures Seizures are a potential risk with atomoxetine. ATOMAKTIN should be introduced with caution in patients with a history of seizure. Discontinuation of atomoxetine should be considered in any patient developing a seizure or if there is an increase in seizure frequency where no other cause is identified.
Growth and development Growth and development should be monitored in children and adolescents during treatment with atomoxetine. Patients requiring long-term therapy should be monitored and consideration should be given to dose reduction or interrupting therapy in children and adolescents who are not growing or gaining weight satisfactorily. Clinical data do not suggest a deleterious effect of atomoxetine on cognition or sexual maturation; however the amount of available long-term data is limited. Therefore, patients requiring long-term therapy should be carefully monitored.
New - onset or worsening of Comorbid Depression, Anxiety and Tics In a controlled study of paediatric patients with ADHD and co morbid chronic motor tics or Tourette's Disorder, atomoxetine-treated patients did not experience worsening of tics compared to placebo-treated patients. In a controlled study of adolescent patients with ADHD and co morbid Major Depressive Disorder, atomoxetine-treated patients did not experience worsening of depression compared to placebo-treated patients. In two controlled studies (one in paediatric patients and one in adult patients) of patients with ADHD and co-morbid anxiety disorders, atomoxetine-treated patients did not experience worsening of anxiety compared to placebo-treated patients. There have been post marketing reports of anxiety and depression or depressed mood and reports of tics in patients taking atomoxetine (see section 4.8).
Patients who are being treated for ADHD with atomoxetine should be monitored for the appearance or worsening of anxiety symptoms, depressed mood and depression or tics.
Effects on micturition In adults ADHD controlled trials, the rates of urinary retention and urinary hesitation were increased among the atomoxetine subjects compared with placebo subjects. A complaint of urinary retention or urinary hesitancy should be considered potentially related to ATOMAKTIN.
Elderly use The safety and efficacy of ATOMAKTIN in elderly patients have not been established.
Paediatric population under six years of age ATOMAKTIN should not be used in patients less than six years of age as efficacy and safety have not been established in this age group.
Special populations ATOMAKTIN has been used in patients with ADHD without deterioration of conditions of motor tics. Tourette syndrome (children), co-morbid major depressive disorder (adolescents) and anxiety disorders (adults).
Other therapeutic use ATOMAKTIN is not indicated for the treatment of major depressive episodes and/or anxiety as the results of clinical trials in adults in these conditions, where ADHD is not present, did not show an effect compared to placebo (see section 5.1).
4.5 Interaction with other medicines and other forms of interaction
Effects of other medicines on ATOMAKTIN MAOIs ATOMAKTIN should not be used with MAOIs (see section 4.3). CYP2D6 inhibitors (SSRIs (e.g. fluoxetine, paroxetine), quinidine, terbinafine) In patients receiving these medicines, atomoxetine exposure may be 6-to 8-fold increased and Css max 3 to 4 times higher, because it is metabolised by the CYP2D6 pathway. Slower titration and final lower dosage of atomoxetine may be necessary in patients who are already taking CYP2D6 inhibitor medicines. If a CYP2D6 inhibitor is prescribed or discontinued after titration to the appropriate atomoxetine dose has occurred, the clinical response and tolerability should be re-evaluated for that patient to determine if dose adjustment is needed. Caution is advised when combining atomoxetine with potent inhibitors of cytochrome P450 enzymes other than CYP2D6 in patients who are poor CYP2D6 metabolisers as the risk of clinically relevant increases in atomoxetine exposure in vivo is unknown.
Salbutamol (or other beta 2 agonists) ATOMAKTIN should be administered with caution to patients treated with high dose nebulised or systemically administered salbutamol (or other beta 2 agonists) because cardiovascular effects can be potentiated. Contradictory findings regarding this interaction were found. Systemically administered salbutamol (600 u03bcg i.v. over 2hrs) in combination with atomoxetine (60 mg twice daily for 5 days) induced increases in heart rate and blood pressure. This effect was most marked after the initial coadministration of salbutamol and atomoxetine but returned towards baseline at the end of 8 hours. However, in a separate study the effects on blood pressure and heart rate of a standard inhaled dose of salbutamol (200 u03bcg) were not increased by the short term coadministration of atomoxetine (80 mg once daily for 5 days) in a study of healthy Asian adults who were extensive atomoxetine metabolisers. Similarly heart rate after multiple inhalations of salbutamol (800 u03bcg) did not differ in the presence or absence of atomoxetine. Attention should be paid to monitoring heart rate and blood pressure, and dose adjustments may be justified for either atomoxetine or salbutamol (or other beta2 agonists) in the event of significant increases in heart rate and blood pressure during coadministration of these medicines.
There is the potential for an increased risk of QT interval prolongation when atomoxetine is administered with other QT-prolonging medicines, (such as neuroleptics, class IA and III anti dysrhythmics, moxifloxacin, erythromycin, methadone, mefloquine, tricyclic antidepressants, lithium or cisapride) medicines that cause electrolyte imbalance (such as thiazide diuretics) and medicines that inhibit CYP2D6. Seizures are a potential risk with atomoxetine. Caution is advised with concomitant use of medicines which are known to lower the seizure threshold (such as tricyclic antidepressants or SSRIs, neuroleptics, phenothiazines or butyrophenone, mefloquine, chloroquine, bupropion or tramadol) (see section 4.4).
In addition, caution is advised when stopping concomitant treatment with benzodiazepines due to potential withdrawal seizures.
Anti-hypertensive medicines ATOMAKTIN should be used cautiously with antihypertensive drugs. Because of a possible increase in blood pressure, ATOMAKTIN may decrease the effectiveness of antihypertensive medicines / medicines used to treat hypertension. Attention should be paid to monitoring of blood pressure and review of treatment of ATOMAKTIN or antihypertensive medicines may be justified in the case of significant changes of blood pressure.
Pressor agents or medicines that increase blood pressure Because of possible increase in effects on blood pressure, ATOMAKTIN should be used cautiously with pressor agents or medications that may increase blood pressure (such as salbutamol). Attention should be paid to monitoring of blood pressure, and review of treatment for either ATOMAKTIN or pressor agents may be justified in the case of significant change in blood pressure.
Medicines that Affect Noradrenaline Drugs that affect noradrenaline should be used cautiously when co-administered with ATOMAKTIN because of the potential for additive or synergistic pharmacological effects. Examples include antidepressants such as imipramine, venlafaxine and mirtazapine, or the decongestants pseudoephedrine or phenylephrine.
Medicines that Affect Gastric pH Medicines that elevate gastric pH (magnesium hydroxide/aluminium hydroxide, omeprazole) had no effect on atomoxetine bioavailability. Medicines Highly Bound to Plasma Protein In vitro drug-displacement studies were conducted with atomoxetine and other highly bound medicines at therapeutic concentrations. Warfarin, acetylsalicylic acid, phenytoin, or diazepam did not affect the binding of atomoxetine to human albumin. Similarly, atomoxetine did not affect the binding of these compounds to human albumin.
4.6 Fertility, pregnancy and lactation
Pregnancy Safety and efficacy have not been demonstrated in pregnancy.
Breastfeeding Women using ATOMAKTIN should not breastfeed their infants.
4.7 Effects on ability to drive and use machines
Atomoxetine, as in ATOMAKTIN has been associated with increased rates of fatigue, somnolence, and dizziness relative to placebo in paediatric and adult patients. Patients should be advised to use caution when driving a car or operating hazardous machinery until they are reasonably certain that their performance is not affected by ATOMAKTIN.
4.8 Undesirable effects
Summary of the safety profile The most frequently reported adverse drug reactions in paediatric patients, during placebo-controlled clinical trials with atomoxetine were headache, abdominal pain and decreased appetite. Abdominal pain and decreased appetite are usually transient. Associated with decreased appetite, some patients experienced growth retardation early in therapy in terms of both weight and height gain. On average, after an initial decrease in weight and height gain, patients treated with atomoxetine recovered to mean weight and height as predicted by group baseline data over the long-term treatment. Nausea, vomiting and somnolence may occur. In both paediatric and adult placebo-controlled trials, patients taking atomoxetine experienced increases in heart rate, systolic and diastolic blood pressure (see section 4.4). Because of its effect on noradrenergic tone, orthostatic hypotension and syncope have been reported in patients taking atomoxetine. Atomoxetine should be used with caution in any condition that may predispose patients to hypotension.
The following table of undesirable effects is based on adverse event reporting and laboratory investigations from clinical trials and post marketing spontaneous reports in children and adolescents:
Tabulated summary of adverse reactions
System organ class Frequent Less frequent Frequency unknown
Metabolism and nutrition disorders Appetite decreased Anorexia (loss of appetite)
Psychiatric disorders Irritability, mood swings, insomnia, agitation, libido decreased, sleep disorder anxiety, depression and depressed mood, tics
Suicide-related events, aggression, hostility, emotional lability, restlessness, psychosis (including hallucinations), orgasm abnormal
Nervous system disorders Headache Somnolence (including sedation), Dizziness, tremor, dysgeusia, Syncope, migraine, hypoaesthesia, Seizure.
4.9 Overdose
Signs and symptoms During post marketing, there have been reports of non-fatal acute and chronic overdoses of atomoxetine alone. The most commonly reported symptoms accompanying acute and chronic overdoses were gastrointestinal symptoms somnolence, dizziness, tremor and abnormal behaviour. Hyperactivity and agitation have also been reported. Signs and symptoms consistent with mild to moderate sympathetic nervous system activation (e.g. tachycardia, blood pressure increased, mydriasis, dry mouth) were also observed and reports of pruritus and rash have been received. Most events were mild to moderate. In some cases of overdose involving atomoxetine, seizures have been reported and very rarely QT-prolongation. There have also been reports of fatal, acute overdoses involving a mixed ingestion of atomoxetine and at least one other medicine.
Management An airway should be established. Activated charcoal may be useful in limiting absorption if the patient presents within 1 hour of ingestion. Monitoring of cardiac and vital signs is recommended, along with appropriate symptomatic and supportive measures. The patient should be observed for a minimum of 6 hours. Because atomoxetine is highly protein-bound, dialysis is not likely to be useful in the treatment of overdose.