Comirnaty Concentrate For Dispersion For 0,3 mL Injection
PDF Leaflet Revision Date: 23 August 2023
Clinical Summary
Quick overview from the medicine insert
Indication
Active immunisation to prevent COVID-19 in individuals 12 years and older.
Dosage (summary)
Primary course: 2 doses (0.3 mL each) IM, 3 weeks apart; booster: 0.3 mL IM.
Special Populations
- Elderly
- Severely immunocompromised
Pregnancy & Breastfeeding
Can be used during pregnancy and breastfeeding; no adverse effects expected.
Contraindications
- Hypersensitivity to vaccine components
Common side effects
- Injection site pain
- Fatigue
- Headache
- Myalgia
- Chills
Counselling Points
- Monitor for 15 mins post-vaccination
- Report any severe reactions immediately
Serious warnings
- Risk of anaphylaxis
- Increased risk of myocarditis and pericarditis
Important Disclaimer
The Comirnaty Concentrate For Dispersion For 0,3 mL Injection professional information leaflet below is the property of Pfizer Laboratories and is provided on Medinsert exactly as issued, with no.. alterations or editorial changes. We make every effort to keep content current by updating documents as soon as new versions become available. Medinsert serves as a trusted access point for healthcare professionals, but does not replace official sources or clinical judgement. For more details, please read our full disclaimer. read more>>
 Healthcare Professionals Only
This content is for registered healthcare professionals
Sign in or create a free account to read the full package insert.
Free for HPCSA-registered professionals. Powered by Medinsert.
Clinical Particulars
Section 4 of the official insert — extracted exactly as issued, no alterations
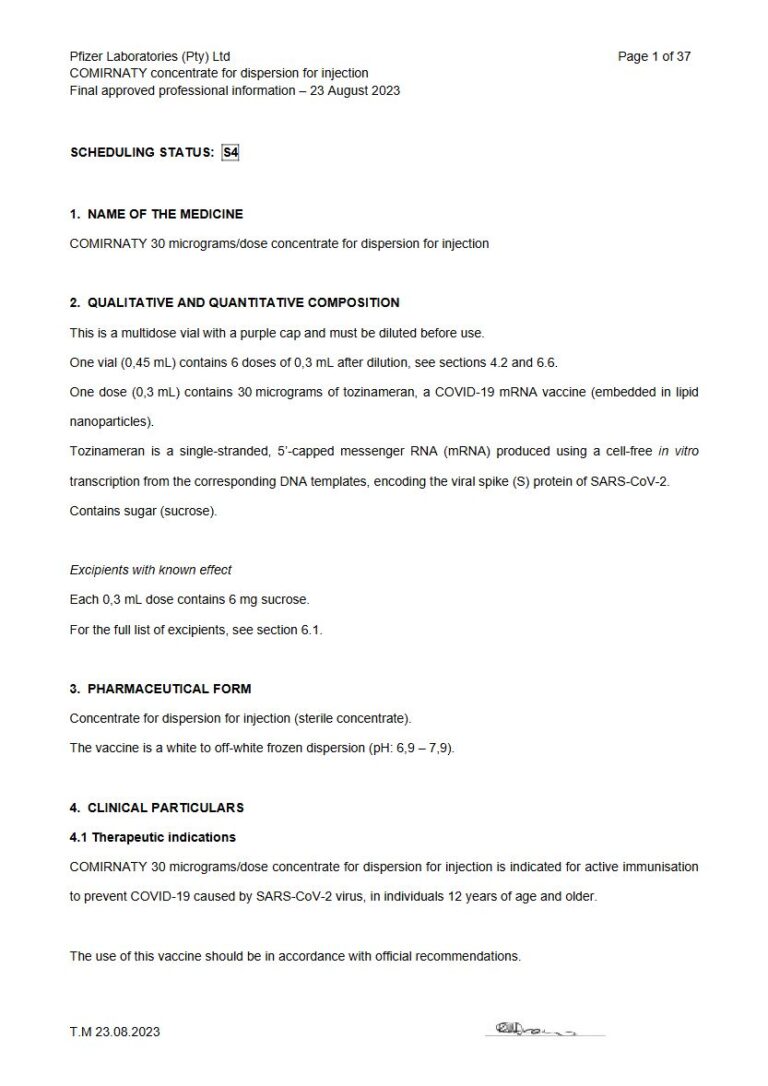
4.1 Therapeutic indications
COMIRNATY 30 micrograms/dose concentrate for dispersion for injection is indicated for active immunisation to prevent COVID-19 caused by SARS-CoV-2 virus, in individuals 12 years of age and older. The use of this vaccine should be in accordance with official recommendations.
4.2 Posology and method of administration
Posology
Primary vaccination course
Individuals 12 years of age and older
COMIRNATY is administered intramuscularly after dilution as a primary course of 2 doses (0,3 mL each). It is recommended to administer the second dose 3 weeks after the first dose (see sections 4.4 and 5.1).
Severely immunocompromised aged 12 years and older
A third primary course dose may be administered intramuscularly at least 28 days after the second dose to individuals who are severely immunocompromised (see section 4.4).
Interchangeability
The interchangeability of COMIRNATY with COVID-19 vaccines from other manufacturers to complete the primary course has not been established. Individuals who have received 1 dose of COMIRNATY should continue to receive COMIRNATY to complete the primary course. Doses of COMIRNATY 30 micrograms/dose concentrate for dispersion for injection after dilution (supplied in a vial with a purple cap) and COMIRNATY READY TO USE ADULT VACCINE 30 micrograms/dose dispersion for injection (supplied in a vial with a grey cap) are considered interchangeable.
Booster dose
The booster dose of COMIRNATY is 0,3 mL given intramuscularly. A booster dose may be given in individuals 12 years of age and older. There should be an interval of at least 3 months between administration of COMIRNATY and the last prior dose of a COVID-19 vaccine.
Special populations
Elderly
No dosage adjustment is required in elderly individuals u2265 65 years of age.
Paediatric population
There is a paediatric formulation available for individuals 5 to 11 years of age (i.e. 5 to less than 12 years of age). For details, please refer to the professional information for COMIRNATY DILUTE TO USE PAEDIATRIC VACCINE 10 micrograms/dose concentrate for dispersion for injection. The safety and efficacy of COMIRNATY in infants aged less than 6 months have not yet been established.
Method of administration
COMIRNATY 30 micrograms/dose concentrate for dispersion for injection should be administered intramuscularly after dilution (see section 6.6). After dilution, vials of COMIRNATY contain six doses of 0,3 mL of vaccine. In order to extract six doses from a single vial, low dead-volume syringes and/or needles should be used. The low dead-volume syringe and needle combination should have a dead volume of no more than 35 microlitres. If standard syringes and needles are used, there may not be sufficient volume to extract a sixth dose from a single vial. Irrespective of the type of syringe and needle:
- Each dose must contain 0,3 mL of vaccine
- If the amount of vaccine remaining in the vial cannot provide a full dose of 0,3 mL, discard the vial and any excess volume
- Do not pool excess vaccine from multiple vials
The preferred site is the deltoid muscle of the upper arm. Do not inject the vaccine intravascularly, subcutaneously or intradermally. The vaccine should not be mixed in the same syringe with any other vaccines or medicines. For precautions to be taken before administering the vaccine, see section 4.4. For instructions regarding thawing, handling and disposal of the vaccine, see section 6.6.
4.3 Contraindications
Hypersensitivity to COVID-19 mRNA vaccine (nucleoside modified) or to any of the excipients listed in section 6.1 [((4-hydroxybutyl)azanediyl)bis(hexane-6,1-diyl)bis(2-hexyldecanoate) (ALC-0315); 2 [(polyethylene glycol)-2000]-N,N-ditetradecylacetamide (ALC-0159); 1,2-Distearoyl-sn-glycero-3-phosphocholine (DSPC); cholesterol; potassium chloride; potassium dihydrogen phosphate sodium chloride; disodium phosphate dihydrate; sucrose; water for injections; sodium hydroxide and hydrochloric acid].
4.4 Special warnings and precautions for use
Traceability
In order to improve the traceability of biological medicines, the name and the batch number of the administered medicine should be clearly recorded.
General recommendations
Hypersensitivity and anaphylaxis
Events of anaphylaxis have been reported. Appropriate medical treatment and supervision should always be readily available in case of an anaphylactic reaction following the administration of the vaccine. Close observation for at least 15 minutes is recommended following vaccination. No further dose of the vaccine should be given to those who have experienced anaphylaxis after a prior dose of COMIRNATY.
Myocarditis and pericarditis
There is an increased risk of myocarditis and pericarditis following vaccination with COMIRNATY. These conditions can develop within just a few days after vaccination and have primarily occurred within 14 days. They have been observed more often after the second vaccination, and more often in younger males. Available data suggest that the course of myocarditis and pericarditis following vaccination is not different from myocarditis or pericarditis in general (see section 4.8). Medical practitioners should be alert to the signs and symptoms of myocarditis and pericarditis. Vaccinees (including parents or caregivers) should be instructed to seek immediate medical attention if they develop symptoms indicative of myocarditis or pericarditis such as (acute and persisting) chest pain, shortness of breath, or palpitations following vaccination. Medical practitioners should consult guidance and/or specialists to diagnose and treat this condition.
Anxiety-related reactions
Anxiety-related reactions, including vasovagal reactions (syncope), hyperventilation or stress u2010 related reactions (e.g. dizziness, palpitations, increases in heart rate, alterations in blood pressure, paraesthesia, hypoaesthesia and sweating) may occur in association with the vaccination process itself. Stress-related reactions are temporary and resolve on their own. Individuals should be advised to bring symptoms to the attention of the vaccination provider for evaluation. It is important that precautions are in place to avoid injury from fainting.
Concurrent illness
Vaccination should be postponed in individuals suffering from acute severe febrile illness or acute infection. The presence of a minor infection and/or low-grade fever should not delay vaccination.
Thrombocytopenia and coagulation disorders
The vaccine should be given with caution in individuals receiving anticoagulant therapy or those with thrombocytopenia or any coagulation disorder (such as haemophilia) because bleeding or bruising may occur following an intramuscular administration in these individuals.
Immunocompromised individuals
The efficacy and safety of COMIRNATY has not been assessed in immunocompromised individuals, including those receiving immunosuppressant therapy. The efficacy of COMIRNATY may be lower in immunocompromised individuals. The recommendation to consider a third dose in severely immunocompromised individuals is based on limited serological evidence from a case series in the literature from the clinical management of patients with iatrogenic immunocompromisation after solid organ transplantation (see section 4.2).
Duration of protection
The duration of protection afforded by the vaccine is unknown as it is still being determined by ongoing clinical trials.
Limitations of vaccine effectiveness
Vaccination with COMIRNATY may not protect all vaccine recipients. Individuals may not be fully protected until 7 days after their second dose of vaccine.
Excipients
COMIRNATY contains less than 1 mmol potassium (39 mg) per dose, that is to say essentially u2018potassium-freeu2019. COMIRNATY contains less than 1 mmol sodium (23 mg) per dose, that is to say essentially u2018sodium-freeu2019. Refer to sections 4.3 or 6.1 for the full list of excipients.
4.5 Interaction with other medicines and other forms of interaction
No interaction studies have been performed. Concomitant administration of COMIRNATY with other vaccines has not been studied.
4.6 Fertility, pregnancy and lactation
Pregnancy
A large amount of observational data from pregnant women vaccinated with COMIRNATY during the second and third trimester have not shown an increase in adverse pregnancy outcomes. While data on pregnancy outcomes following vaccination during the first trimester are presently limited, no increased risk for miscarriage has been seen. Animal studies do not indicate direct or indirect harmful effects with respect to pregnancy, embryo/foetal development, parturition or post-natal development (see section 5.3). COMIRNATY can be used during pregnancy.
Breastfeeding
No effects on the breastfed newborn/infant are anticipated since the systemic exposure of breastfeeding woman to COMIRNATY is negligible. Observational data from women who were breastfeeding after vaccination have not shown a risk for adverse effects in breastfed newborns/infants. COMIRNATY can be used during breastfeeding.
Fertility
Animal studies do not indicate direct or indirect harmful effects with respect to reproductive toxicity (see section 5.3).
4.7 Effects on ability to drive and use machines
COMIRNATY has no or negligible influence on the ability to drive and use machines. However, some of the effects mentioned under section 4.8 may temporarily affect the ability to drive or use machines.
4.8 Undesirable effects
Summary of the safety profile
Participants 16 years of age and older - after 2 doses
In Study 2, a total of 22 026 participants 16 years of age or older received at least 1 dose of COMIRNATY and a total of 22 021 participants 16 years of age or older received placebo (including 138 and 145 adolescents 16 and 17 years of age in the vaccine and placebo groups, respectively). A total of 20 519 participants 16 years of age or older received 2 doses of COMIRNATY. At the time of the analysis of Study 2 with a data cut-off of 13 March 2021 for the placebo-controlled blinded follow-up period up to the participantsu2019 unblinding dates, a total of 25 651 (58,2 %) participants (13 031 COMIRNATY and 12 620 placebo) 16 years of age and older were followed up for u2265 4 months after the second dose. This included a total of 15 111 (7 704 COMIRNATY and 7 407 placebo) participants 16 to 55 years of age and a total of 10 540 (5 327 COMIRNATY and 5 213 placebo) participants 56 years and older.
The most frequent adverse reactions in participants 16 years of age and older that received 2 doses were injection site pain (> 80 %), fatigue (> 60 %), headache (> 50 %), myalgia (> 40 %), chills (> 30 %), arthralgia (> 20 %), pyrexia and injection site swelling (> 10 %) and were usually mild or moderate in intensity and resolved within a few days after vaccination. A slightly lower frequency of reactogenicity events was associated with greater age.
The safety profile in 545 participants 16 years of age and older receiving COMIRNATY, that were seropositive for SARS-CoV-2 at baseline, was similar to that seen in the general population.
Adolescents 12 to 15 years of age u2013 after 2 doses
In an analysis of long-term safety follow-up in Study 2, 2 260 adolescents (1 131 COMIRNATY and 1 129 placebo) were 12 to 15 years of age. Of these, 1 559 adolescents (786 COMIRNATY and 773 placebo) have been followed for u2265 4 months after the second dose of COMIRNATY. The safety evaluation in Study 2 is ongoing. The overall safety profile of COMIRNATY in adolescents 12 to 15 years of age was similar to that seen in participants 16 years of age and older. The most frequent adverse reactions in adolescents 12 to 15 years of age that received 2 doses were injection site pain (> 90 %), fatigue and headache (> 70 %), myalgia and chills (> 40 %), arthralgia and pyrexia (> 20 %).
Participants 12 years of age and older u2013 after booster dose
The safety of a booster dose of COMIRNATY in participants 12 years of age and older is inferred from safety data from studies of a booster dose of COMIRNATY in participants 16 years of age and older. A subset from Study 2 Phase 2/3 participants of 306 adults 18 to 55 years of age who completed the original COMIRNATY 2-dose course, received a booster dose of COMIRNATY approximately 6 months (range of 4,8 to 8,0 months) after receiving Dose 2. Overall, participants who received a booster dose, had a median follow-up time of 8,3 months (range 1,1 to 8,5 months) and 301 participants had been followed for u2265 6 months after the booster dose to the cut-off date (22 November 2021). The overall safety profile for the booster dose was similar to that seen after 2 doses. The most frequent adverse reactions in participants 18 to 55 years of age were injection site pain (> 80 %), fatigue (> 60 %), headache (> 40 %), myalgia (> 30 %), chills and arthralgia (> 20 %).
In Study 4, a placebo-controlled booster study, participants 16 years of age and older recruited from Study 2 received a booster dose of COMIRNATY (5 081 participants), or placebo (5 044 participants) at least 6 months after the second dose of COMIRNATY. Overall, participants who received a booster dose, had a median follow-up time of 2,58 months (range 0,3 to 7,5 months) after the booster dose in the blinded placebo-controlled follow-up period to the cut-off date (8 February 2022). Of these, 1 281 participants (895 COMIRNATY and 386 placebo) have been followed for u2265 4 months after the booster dose of COMIRNATY. No new adverse reactions of COMIRNATY were identified.
Participants 12 years of age and older u2013 after subsequent booster doses
The safety of a booster dose of COMIRNATY in participants 12 years of age and older is inferred from safety data from studies of a booster dose of COMIRNATY in participants 18 years of age and older. A subset of 325 adults 18 to u2264 55 years of age who had completed 3 doses of COMIRNATY, received a booster (fourth dose) of COMIRNATY 90 to 180 days after receiving Dose 3. Participants who received a booster (fourth dose) of COMIRNATY had a median follow-up time of 1,4 months up to a data cut-off date of 11 March 2022. The most frequent adverse reactions in these participants were injection site pain (> 70 %), fatigue (> 60 %), headache (> 40 %), myalgia and chills (> 20 %), and arthralgia (> 10 %). In a subset from Study 4 (Phase 3), 305 adults > 55 years of age who had completed 3 doses of COMIRNATY, received a booster (fourth dose) of COMIRNATY 5 to 12 months after receiving Dose 3. Participants who received a booster (fourth dose) of COMIRNATY had a median follow-up time of at least 1,7 months up to a data cut-off date of 16 May 2022. The overall safety profile for the COMIRNATY booster (fourth dose) was similar to that seen after the COMIRNATY booster (third dose). The most frequent adverse reactions in participants > 55 years of age were injection site pain (> 60 %), fatigue (> 40 %), headache (> 20 %), myalgia and chills (> 10 %).
Booster dose following primary vaccination with another authorised COVID-19 vaccine
In 5 independent studies on the use of a COMIRNATY booster dose in individuals who had completed primary vaccination with another authorized COVID-19 vaccine (heterologous booster dose), no new safety issues were identified (see section 5.1).
Tabulated list of adverse reactions from clinical studies and post-authorisation in individuals 12 years of age and older
Adverse reactions observed during clinical studies are listed below according to the following frequency categories: Very common (u2265 1/10), common (u2265 1/100 to < 1/10), uncommon (u2265 1/1 000 to < 1/100), rare (u2265 1/10 000 to < 1/1 000), very rare (< 1/10 000), not known (cannot be estimated from the available data).
Table 1: Adverse reactions from COMIRNATY clinical trials in individuals 12 years of age and older
System organ class Frequency Adverse reaction Blood and lymphatic system disorders Uncommon Lymphadenopathy
4.9 Overdose
Overdose data is available from 52 study participants included in the clinical trial that due to an error in dilution received 58 micrograms of COMIRNATY. The vaccine recipients did not report an increase in reactogenicity or adverse reactions. In the event of overdose, monitoring of vital functions and possible symptomatic treatment is recommended.